Close

UK Exchequer Wants More Pharma Investment Coming in Britain

Note* - All images used are for editorial and illustrative purposes only and may not originate from the original news provider or associated company.

Related stories

Managing Quality and Regulatory Risks in Pharmaceutical Technology Transfer

Technology transfer moves more than manufacturing instructions. It moves process knowledge, supplier dependencies, labeling requirements, packaging specifications, and the evidence needed to demonstrate control. For medical device manufacturers, including those supporting drug-device combination products, these interfaces can become recurring sources of compliance gaps during site transitions, acquisitions, and remediation. The central problem is often not a missing procedure. It is a quality system that allows connected activities to change independently. A label revision may be approved while packaging artwork, supplier instructions, or production-system data still reflect an obsolete version. Staying inspection-ready requires manufacturers to manage these dependencies as one controlled system. Why Compliance Gaps Keep Recurring Findings are treated as documentation defects Revising a procedure can resolve an immediate discrepancy without changing the behavior that caused it. If unclear decision rights, weak escalation, or ineffective training remain, the same failure may surface elsewhere. Corrective and preventive action should distinguish the observed finding from its underlying cause and establish how effectiveness will be demonstrated. Ownership is divided across connected functions Quality, Regulatory Affairs, labeling, packaging engineering, operations, suppliers, and digital-system owners often maintain separate workflows. Each team may complete its assigned task while an interface remains uncontrolled. For example, changing an instruction for use can affect packaging configuration, translations, technical documentation, inventory disposition, and release controls. Transitions expose unresolved weaknesses Acquisitions and site transfers can introduce conflicting procedures, incomplete knowledge transfer, or different assumptions about validated processes. Regulatory changes add further demands. When these events coincide with CAPA or change-control backlogs, teams may prioritize administrative closure over risk reduction. Without a coordinated impact assessment, local fixes can create new gaps downstream. Establish the Regulatory Baseline Inspection readiness starts with knowing which requirements apply to the product, activity, and market. Pharmaceutical, device, and combination-product obligations are not interchangeable; transfer planning should identify the applicable requirements rather than assume one quality framework covers every operation. According to FDA’s Quality Management System Regulation overview and FAQs, the QMSR became effective on February 2, 2026, incorporates ISO 13485:2016 by reference, and introduced the updated device inspection process under Compliance Program 7382.850. ISO 13485 certification does not exempt a manufacturer from FDA inspection or guarantee compliance with FDA requirements. For devices marketed in the European Union, Regulation (EU) 2017/745, Annex I, Section 23, provides the primary requirements for information supplied with devices, including labels and instructions for use. Teams should assess the applicable provisions alongside relevant language, product-specific, and market requirements. A labeling review should therefore evaluate required content and consistency with the device’s intended purpose and supporting documentation—not merely artwork accuracy. How to Identify the Real Gaps Begin with a cross-functional assessment that follows actual product and information flows. Select representative transfers, label changes, supplier changes, and CAPAs, then trace each from initiation through implementation and effectiveness review. Map dependencies: Identify affected specifications, suppliers, packaging components, systems, validation activities, and training. Compare records with practice: Check whether approved procedures match what operators and reviewers actually do. Test traceability: Follow a requirement through technical documentation, labeling, production instructions, and release evidence. Examine recurrence: Look for repeated deviations, reopened actions, overdue changes, and findings shared across sites. Assess capacity: Determine whether qualified owners have enough time and authority to complete corrective work. Prioritize gaps by potential patient or user harm, regulatory impact, likelihood of recurrence, and detectability. Document the rationale; an old overdue item is not automatically more important than a newly identified high-risk weakness. Align Quality, Labeling, Packaging, and Change Control Create a coordinated remediation plan with one accountable owner for each action, named contributors, acceptance criteria, and required evidence. Separate immediate containment from systemic correction. Containment might prevent obsolete labels from being issued; correction might redesign approval and implementation controls so obsolete versions cannot return. Use change control as the integration point. Before approval, evaluate effects on regulatory submissions or notifications, technical documentation, supplier controls, packaging performance, validation, electronic master data, inventory, and training. Record both identified impacts and justified decisions that particular activities are unaffected. During technology transfer, define readiness gates before production or release responsibilities move. These should address the transfer of process knowledge, acceptance criteria, receiving-site capability, required qualification or validation, approved documents, and personnel competence. The appropriate evidence depends on the product and process; a generic checklist cannot replace a documented assessment. Finally, verify implementation where the work happens. Confirm that current instructions are available, suppliers received approved requirements, systems contain correct versions, and affected personnel can perform their tasks. A signed approval is not evidence that every downstream change took effect. Mistakes That Undermine Inspection Readiness Closing CAPAs on task completion alone: Effectiveness checks should test whether the underlying failure has been controlled. Updating labels in isolation: Assess packaging, supplier, system, documentation, and inventory consequences. Counting closures instead of evidence: Dashboards need defensible records behind their status indicators. ...

State of API & Peptide CDMO Capacity

Pharmaceutical companies planning peptide or complex small-molecule launches are...

What Goes Into Synthetic Urine? The Chemistry Behind Replicating a Biological Fluid

Synthetic urine sounds simple until you look closely at...
- Advertisement -

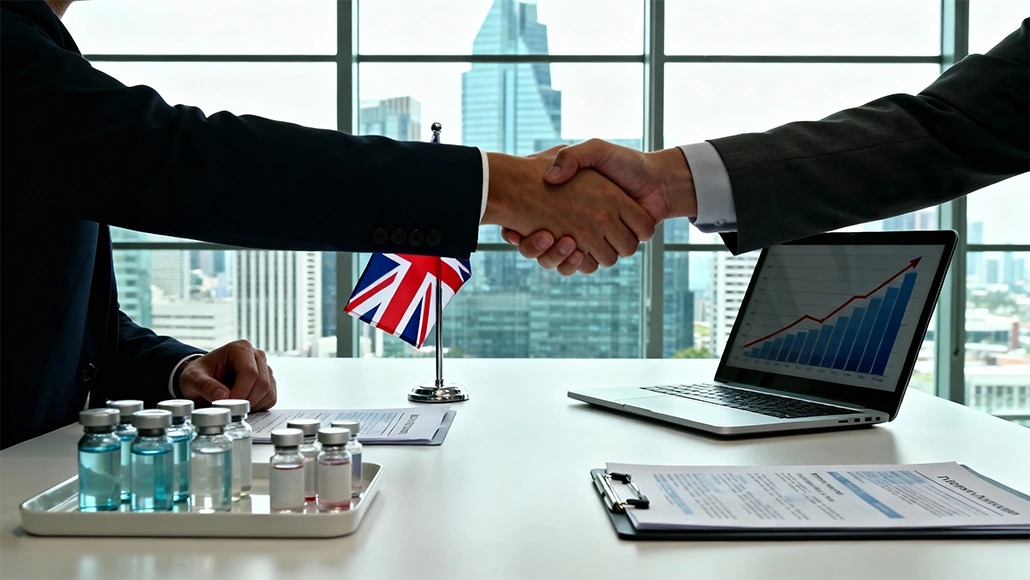
Rachel Reeves, the UK Chancellor of the Exchequer, has asked more pharma investment coming in Britain, even as she doubles down on the commitment by the government when it comes to competitive drug pricing.

Speaking to the reporters on the sidelines of the international meetings in Washington, Reeves went ahead and struck a delicate balance by positioning the UK to be an irresistible hub for life sciences innovation while at the same time, firmly rejecting any sort of a rollback on efforts so as to keep medicines affordable for the NHS.

According to Reeves, they do need to make sure that it happens to be a very attractive place for pharmaceuticals, and that includes pricing too. However, in return for that, they want to see more pharma investment coming in Britain. Reeves’ words come at a very precarious moment for the industry, as there is a string of high-profile pullbacks by certain major drugmakers that have cast a shadow over the ambitions of the UK to become the premier life sciences powerhouse of Europe by 2030.

A Sector Which Is Under Strain

It is well to be noted that the pharmaceutical industry has long been regarded as the crown jewels of Britain, as it contributes more than £108 billion a year to the economy and also supports hundreds of thousands of jobs. Right from the hallowed labs of Oxford and Cambridge to the sprawling manufacturing sites based in the North, the UK happens to boast of a world-class ecosystem when it comes to drug discovery and development. Still, this year alone, the sector has gone on to witness almost £2 billion when it comes to planned investments that have been either scrapped or paused, as per the industry estimates.

The fact remains that the casualties are stark. In September 2025, Merck, the US giant that is known as MSD outside of the US, abandoned a £1 billion research hub in King’s Cross in London, thereby axing 125 scientific positions and also citing a quite unattractive business environment. Eli Lilly followed thereafter by freezing many of the UK projects, whereas AstraZeneca, which is a British-Swedish stalwart, halted a £200 million R&D expansion at Cambridge and, earlier in 2025, also ditched a £450 million vaccine plant based out of Merseyside. These decisions, as per the executives, stem from a very toxic mix when it comes to regulatory hurdles, which is indeed a very controversial clawback tax as far as the drug sales are concerned, and also the strict NHS reimbursement rules, which happen to prioritize value-for-money over incentives for innovation.

The chief executive of the Association of the British Pharmaceutical Industry (ABPI), Richard Torbett, has gone ahead and described the trend as a chronic underinvestment that warns it inflicts suffering onto patients by delaying the access to cutting-edge treatments. This indeed has to be a wake-up call, said Torbett to BBC Radio 4. The UK really happens to have the potential to be winning more billion-pound R&D investments, and rather the fact of the matter is that they are losing them.

Lord Patrick Vallance, the science minister and also a former GlaxoSmithKline executive, echoed these concerns, acknowledging the fact that declining NHS spending on medicines, down in real terms in recent years, goes on to risk eroding the competitive edge of the UK. It is worth noting that Vallance is set to join industry leaders, which includes the likes of UK president of AstraZeneca, Tom Keith-Roach, for high-level talks that are aimed at salvaging the momentum of the sector.

Elevating Clinical Trials and Streamlining Approvals

At the center of this impasse happens to be a transatlantic tussle on the drug costs. The UK has long negotiated its lower prices for medicines by way of its NHS framework, which is a model that has already saved billions; however, it has irked American policymakers who still argue that it amounts to freeloading on US innovation. Apparently, the administration of President Donald has amplified this rhetoric by way of threatening 100% tariffs when it comes to pharma imports unless the firms relocate their manufacturing to America, which is a move that could ripple quite painfully via global supply chains.

Interestingly, tensions boiled over at a private dinner in September 2025 between Reeves and Warren Stephens, the US Ambassador to the UK, where the envoy reportedly asked the chancellor to sweeten the pricing deals or face an investment exodus. Trump has gone on to float the most favored nation policy, which could as well cap the US drug prices at their lowest levels that are paid abroad, therefore potentially forcing Britain to push up its own rates or else watch American markets turn dry for the UK exporters.

But Reeves is unyielding. Her government has already offered what it calls a generous as well as unprecedented £1 billion package for three years so as to bolster clinical trials and also streamline approvals, therefore cutting review times to less than 150 days. Still, the pharma leaders remain a bit skeptical, pointing to much broader regulatory burdens and also an uncertain post-Brexit spectrum as deterrents.

Luring the FDI to Improve the Pharmaceutical Sector

The intervention by Rachel Reeves is more than a sector-specific plea – it happens to be a microcosm of a high-stakes economic reset by Labour under Keir Starmer, the UK Prime Minister. Since taking office in July 2024, the chancellor, who, by the way, is the first woman in the role, has indeed championed securonomics, which is a blend of fiscal discipline and private-sector unlocking that’s designed to roll out the fastest G7 growth.

With public finances getting strained and the taxes off-limits for hikes when it comes to corporations or income, her strategy happens to hinge on luring the foreign direct investment, pharma included, by way of stability and targeted reforms.

With regard to this, Reeves has already gone on to ease the banking separations, pushed pension funds towards much riskier UK assets, and even floated a sovereign health fund so as to inject state cash into drug development. She has also culled the competition watchdog’s leadership so as to signal a lighter regulatory touch. However, with Wall Street at a high amid tariff threats from Trump and China’s rare earth export curbs also adding to the global volatility, the pitch by the chancellor feels like a high-wire act.

Critics, which includes the opposition voices on social media, decry pharma retreats as a brutal £1bn blow when it comes to growth plans, with one user saying as to how much more of Rachel Reeves can Britain actually take. Industry watchers see a vicious cycle – which is less investment means slower innovation and higher long-term NHS costs, along with a dimmer allure for projects that lie ahead.

Doing the Balancing Act

It is well to be noted that the deeper dilemma happens to be ethical as much as economic. As one analyst framed it on social media, it’s a question of who deserves to live longer, and that too at what price? The fact is that affordable drugs save lives and stretch the public budgets; however, without strong R&D funding, the pipeline of breakthroughs, from cancer therapies to next-gen vaccines, would run dry.

Notably, the remarks made by Reeves in Washington signal a very practical pivot – the concessions on attractiveness, but no surrender when it comes to affordability. Whether this sort of quid pro quo sways the boardrooms or not remains to be seen. As the chancellor gets back to London ahead of the autumn budget, the focus is on her to go ahead and deliver wins that are tangible– or risk the cold shoulder by pharma turning in a full freeze-out.

As for now, the life sciences dream happens to hang in the balance, which is indeed a testament to the bold vision of the chancellor and the prominent headwinds that it faces.
Never miss a pharmaceutical headline

The pharmaceutical industry moves fast – stay on top of it with our must - read briefings.

  • The top pharma and life sciences stories, straight to your inbox
  • The biggest news, features, interviews, and analysis
  • Dedicated coverage of the key developments driving the global pharmaceutical sector

Subscribe

- Never miss a story with notifications

- Gain full access to our premium content

- Browse free from any location or device.

Media Packs

Expand Your Reach With Our Customized Solutions Empowering Your Campaigns To Maximize Your Reach & Drive Real Results!

– Access the Media Pack Now

– Book a Conference Call

– Leave Message for Us to Get Back

Latest stories

Related stories

Managing Quality and Regulatory Risks in Pharmaceutical Technology Transfer

Technology transfer moves more than manufacturing instructions. It moves process knowledge, supplier dependencies, labeling requirements, packaging specifications, and the evidence needed to demonstrate control. For medical device manufacturers, including those supporting drug-device combination products, these interfaces can become recurring sources of compliance gaps during site transitions, acquisitions, and remediation. The central problem is often not a missing procedure. It is a quality system that allows connected activities to change independently. A label revision may be approved while packaging artwork, supplier instructions, or production-system data still reflect an obsolete version. Staying inspection-ready requires manufacturers to manage these dependencies as one controlled system. Why Compliance Gaps Keep Recurring Findings are treated as documentation defects Revising a procedure can resolve an immediate discrepancy without changing the behavior that caused it. If unclear decision rights, weak escalation, or ineffective training remain, the same failure may surface elsewhere. Corrective and preventive action should distinguish the observed finding from its underlying cause and establish how effectiveness will be demonstrated. Ownership is divided across connected functions Quality, Regulatory Affairs, labeling, packaging engineering, operations, suppliers, and digital-system owners often maintain separate workflows. Each team may complete its assigned task while an interface remains uncontrolled. For example, changing an instruction for use can affect packaging configuration, translations, technical documentation, inventory disposition, and release controls. Transitions expose unresolved weaknesses Acquisitions and site transfers can introduce conflicting procedures, incomplete knowledge transfer, or different assumptions about validated processes. Regulatory changes add further demands. When these events coincide with CAPA or change-control backlogs, teams may prioritize administrative closure over risk reduction. Without a coordinated impact assessment, local fixes can create new gaps downstream. Establish the Regulatory Baseline Inspection readiness starts with knowing which requirements apply to the product, activity, and market. Pharmaceutical, device, and combination-product obligations are not interchangeable; transfer planning should identify the applicable requirements rather than assume one quality framework covers every operation. According to FDA’s Quality Management System Regulation overview and FAQs, the QMSR became effective on February 2, 2026, incorporates ISO 13485:2016 by reference, and introduced the updated device inspection process under Compliance Program 7382.850. ISO 13485 certification does not exempt a manufacturer from FDA inspection or guarantee compliance with FDA requirements. For devices marketed in the European Union, Regulation (EU) 2017/745, Annex I, Section 23, provides the primary requirements for information supplied with devices, including labels and instructions for use. Teams should assess the applicable provisions alongside relevant language, product-specific, and market requirements. A labeling review should therefore evaluate required content and consistency with the device’s intended purpose and supporting documentation—not merely artwork accuracy. How to Identify the Real Gaps Begin with a cross-functional assessment that follows actual product and information flows. Select representative transfers, label changes, supplier changes, and CAPAs, then trace each from initiation through implementation and effectiveness review. Map dependencies: Identify affected specifications, suppliers, packaging components, systems, validation activities, and training. Compare records with practice: Check whether approved procedures match what operators and reviewers actually do. Test traceability: Follow a requirement through technical documentation, labeling, production instructions, and release evidence. Examine recurrence: Look for repeated deviations, reopened actions, overdue changes, and findings shared across sites. Assess capacity: Determine whether qualified owners have enough time and authority to complete corrective work. Prioritize gaps by potential patient or user harm, regulatory impact, likelihood of recurrence, and detectability. Document the rationale; an old overdue item is not automatically more important than a newly identified high-risk weakness. Align Quality, Labeling, Packaging, and Change Control Create a coordinated remediation plan with one accountable owner for each action, named contributors, acceptance criteria, and required evidence. Separate immediate containment from systemic correction. Containment might prevent obsolete labels from being issued; correction might redesign approval and implementation controls so obsolete versions cannot return. Use change control as the integration point. Before approval, evaluate effects on regulatory submissions or notifications, technical documentation, supplier controls, packaging performance, validation, electronic master data, inventory, and training. Record both identified impacts and justified decisions that particular activities are unaffected. During technology transfer, define readiness gates before production or release responsibilities move. These should address the transfer of process knowledge, acceptance criteria, receiving-site capability, required qualification or validation, approved documents, and personnel competence. The appropriate evidence depends on the product and process; a generic checklist cannot replace a documented assessment. Finally, verify implementation where the work happens. Confirm that current instructions are available, suppliers received approved requirements, systems contain correct versions, and affected personnel can perform their tasks. A signed approval is not evidence that every downstream change took effect. Mistakes That Undermine Inspection Readiness Closing CAPAs on task completion alone: Effectiveness checks should test whether the underlying failure has been controlled. Updating labels in isolation: Assess packaging, supplier, system, documentation, and inventory consequences. Counting closures instead of evidence: Dashboards need defensible records behind their status indicators. ...

State of API & Peptide CDMO Capacity

Pharmaceutical companies planning peptide or complex small-molecule launches are...

What Goes Into Synthetic Urine? The Chemistry Behind Replicating a Biological Fluid

Synthetic urine sounds simple until you look closely at...

Danaher Prepares to Introduce Advanced AI-Powered Laboratory

In a significant step for pharmaceutical research, Danaher has...

Subscribe

- Never miss a story with notifications

- Gain full access to our premium content

- Browse free from any location or device.

Media Packs

Expand Your Reach With Our Customized Solutions Empowering Your Campaigns To Maximize Your Reach & Drive Real Results!

– Access theMedia Pack Now

– Book a Conference Call

– Leave Message for Us to Get Back

Translate »