America’s AI Action Plan Boosts Pharmaceutical Innovation

Note* - All images used are for editorial and illustrative purposes only and may not originate from the original news provider or associated company.

Related stories

Billerud Introduces New Pharmaceutical Packaging Option

Billerud has officially introduced PharmaKraft, a specialized fibre-based packaging...

AstraZeneca Advances AI Drug Discovery Deal with insitro

AstraZeneca has inked a billion-dollar AI drug discovery partnership...

Managing Quality and Regulatory Risks in Pharmaceutical Technology Transfer

Technology transfer moves more than manufacturing instructions. It moves process knowledge, supplier dependencies, labeling requirements, packaging specifications, and the evidence needed to demonstrate control. For medical device manufacturers, including those supporting drug-device combination products, these interfaces can become recurring sources of compliance gaps during site transitions, acquisitions, and remediation. The central problem is often not a missing procedure. It is a quality system that allows connected activities to change independently. A label revision may be approved while packaging artwork, supplier instructions, or production-system data still reflect an obsolete version. Staying inspection-ready requires manufacturers to manage these dependencies as one controlled system. Why Compliance Gaps Keep Recurring Findings are treated as documentation defects Revising a procedure can resolve an immediate discrepancy without changing the behavior that caused it. If unclear decision rights, weak escalation, or ineffective training remain, the same failure may surface elsewhere. Corrective and preventive action should distinguish the observed finding from its underlying cause and establish how effectiveness will be demonstrated. Ownership is divided across connected functions Quality, Regulatory Affairs, labeling, packaging engineering, operations, suppliers, and digital-system owners often maintain separate workflows. Each team may complete its assigned task while an interface remains uncontrolled. For example, changing an instruction for use can affect packaging configuration, translations, technical documentation, inventory disposition, and release controls. Transitions expose unresolved weaknesses Acquisitions and site transfers can introduce conflicting procedures, incomplete knowledge transfer, or different assumptions about validated processes. Regulatory changes add further demands. When these events coincide with CAPA or change-control backlogs, teams may prioritize administrative closure over risk reduction. Without a coordinated impact assessment, local fixes can create new gaps downstream. Establish the Regulatory Baseline Inspection readiness starts with knowing which requirements apply to the product, activity, and market. Pharmaceutical, device, and combination-product obligations are not interchangeable; transfer planning should identify the applicable requirements rather than assume one quality framework covers every operation. According to FDA’s Quality Management System Regulation overview and FAQs, the QMSR became effective on February 2, 2026, incorporates ISO 13485:2016 by reference, and introduced the updated device inspection process under Compliance Program 7382.850. ISO 13485 certification does not exempt a manufacturer from FDA inspection or guarantee compliance with FDA requirements. For devices marketed in the European Union, Regulation (EU) 2017/745, Annex I, Section 23, provides the primary requirements for information supplied with devices, including labels and instructions for use. Teams should assess the applicable provisions alongside relevant language, product-specific, and market requirements. A labeling review should therefore evaluate required content and consistency with the device’s intended purpose and supporting documentation—not merely artwork accuracy. How to Identify the Real Gaps Begin with a cross-functional assessment that follows actual product and information flows. Select representative transfers, label changes, supplier changes, and CAPAs, then trace each from initiation through implementation and effectiveness review. Map dependencies: Identify affected specifications, suppliers, packaging components, systems, validation activities, and training. Compare records with practice: Check whether approved procedures match what operators and reviewers actually do. Test traceability: Follow a requirement through technical documentation, labeling, production instructions, and release evidence. Examine recurrence: Look for repeated deviations, reopened actions, overdue changes, and findings shared across sites. Assess capacity: Determine whether qualified owners have enough time and authority to complete corrective work. Prioritize gaps by potential patient or user harm, regulatory impact, likelihood of recurrence, and detectability. Document the rationale; an old overdue item is not automatically more important than a newly identified high-risk weakness. Align Quality, Labeling, Packaging, and Change Control Create a coordinated remediation plan with one accountable owner for each action, named contributors, acceptance criteria, and required evidence. Separate immediate containment from systemic correction. Containment might prevent obsolete labels from being issued; correction might redesign approval and implementation controls so obsolete versions cannot return. Use change control as the integration point. Before approval, evaluate effects on regulatory submissions or notifications, technical documentation, supplier controls, packaging performance, validation, electronic master data, inventory, and training. Record both identified impacts and justified decisions that particular activities are unaffected. During technology transfer, define readiness gates before production or release responsibilities move. These should address the transfer of process knowledge, acceptance criteria, receiving-site capability, required qualification or validation, approved documents, and personnel competence. The appropriate evidence depends on the product and process; a generic checklist cannot replace a documented assessment. Finally, verify implementation where the work happens. Confirm that current instructions are available, suppliers received approved requirements, systems contain correct versions, and affected personnel can perform their tasks. A signed approval is not evidence that every downstream change took effect. Mistakes That Undermine Inspection Readiness Closing CAPAs on task completion alone: Effectiveness checks should test whether the underlying failure has been controlled. Updating labels in isolation: Assess packaging, supplier, system, documentation, and inventory consequences. Counting closures instead of evidence: Dashboards need defensible records behind their status indicators. ...
- Advertisement -

Accelerating Pharmaceutical and Biopharmaceutical Innovation through America’s AI Action Plan

America’s AI Action Plan, unveiled in July 2025, lays out a comprehensive strategy to propel the United States to the forefront of global AI leadership. Within its three pillars—innovation, infrastructure, and international diplomacy—the Plan contains transformative policies tailored to the pharmaceutical and biopharmaceutical industries. By harnessing AI-driven breakthroughs in drug discovery, manufacturing, data sharing, workforce development, and biosecurity, the Plan charts a path toward faster, safer, and more cost-effective development of new therapies.

AI-Enabled Drug Discovery and Automated Laboratories

Central to the Plan’s vision is the use of AI to revolutionise early-stage research. Traditional drug discovery, often a years-long, high-cost endeavour, will be dramatically accelerated by AI tools capable of:

  • Predicting protein structures and binding interactions with unprecedented speed and accuracy
  • Designing candidate molecules through generative models trained on vast chemical libraries
  • Prioritizing compounds most likely to succeed in preclinical testing

To translate these AI predictions into real-world breakthroughs, the Plan directs major investments in automated cloud-enabled laboratories. These next-generation facilities—built through public-private partnerships involving the National Science Foundation, the Department of Energy, and leading biopharmaceutical firms—will integrate robotic experimentation platforms, high-throughput screening, and AI-driven hypothesis generation. By closing the loop between in silico design and in vitro validation, these “lab clouds” aim to reduce the time from target identification to candidate selection from years to months.

Building High-Quality Biomedical Datasets
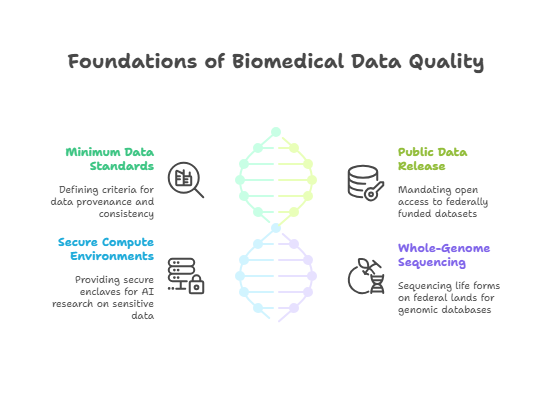
Machine learning models thrive on data. Yet the biopharmaceutical sector’s data landscape has long been fragmented across proprietary silos. The AI Action Plan tackles this bottleneck by:

  1. Establishing Minimum Data Quality Standards
    A national committee will define criteria for data provenance, annotation consistency, and interoperability across modalities—genomic, proteomic, clinical trial, and real-world evidence.
  2. Mandating Public Release of Federally Funded Datasets
    Investigators receiving NIH and National Science Foundation grants must publish non-sensitive, de-identified datasets in open repositories, ensuring broader access for AI model training.
  3. Creating Secure Compute Environments
    To protect proprietary or sensitive data, the Plan funds secure enclaves where researchers can run AI algorithms on restricted datasets without direct data export.
  4. Whole-Genome Sequencing on Federal Lands
    A bold initiative to sequence all life forms on public lands will generate massive genomic databases, fuelling next-generation biopharma AI models for target discovery and biomarker identification.

By weaving these efforts together, the Plan envisions a federated data ecosystem where academic laboratories, biotech startups, and large pharma can collaborate on shared AI challenges while safeguarding patient privacy and intellectual property.

Biosecurity and Safe Biological Design
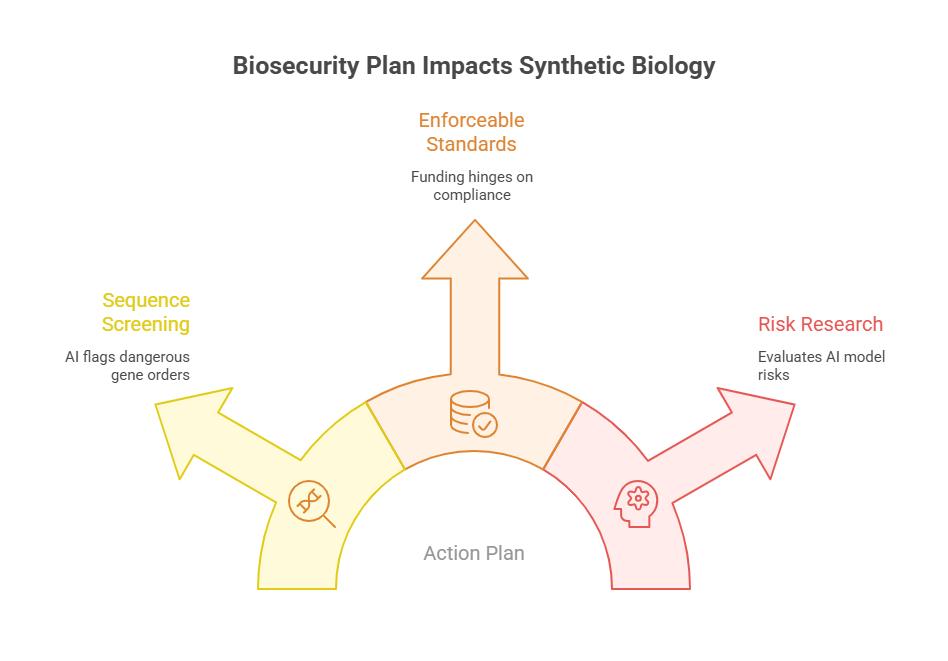
 

AI’s power to design novel genetic sequences raises both medical promise and biosecurity concerns. The Action Plan introduces strict controls on synthetic biology tools used in drug manufacture and vaccine development:

  • Mandatory Sequence Screening
    All companies offering gene synthesis services must deploy robust AI-based screening to flag orders that resemble dangerous pathogens or toxin genes. Real-time alerts and data sharing among providers will prevent malicious misuse.
  • Enforceable Standards
    Instead of voluntary guidelines, federal funding now hinges on compliance with detailed biosecurity protocols. Organisations failing to meet these will lose access to grants and procurement opportunities.
  • Research on Frontier Models’ Risks
    Through NIST and the Department of Commerce, the Plan mandates evaluations of emerging AI models for their potential to generate harmful biological designs, with findings informing regulation.

These measures ensure that as the biopharma sector accelerates vaccine and therapeutic development through AI, it does so within a fortified safety framework.

Next-Generation Biopharmaceutical Manufacturing

Manufacturing biologics—antibodies, cell therapies, and complex proteins—requires stringent process controls and continuous innovation. The AI Action Plan directs targeted funding to integrate AI for:

  • Process Optimization
    Machine learning models will analyse real-time bioreactor sensor data to adjust parameters—pH, temperature, nutrient feed—maximising yield and consistency.
  • Predictive Maintenance
    AI-driven monitoring of equipment health will preemptively flag maintenance needs, reducing downtime and production delays.
  • Quality Control Automation
    Computer vision systems will inspect vials and final product batches for visual defects, enhancing release testing speed and reliability.

Support comes through expanded Small Business Innovation Research grants and Title III Defense Production Act authorities, allowing rapid scale-up of modular, AI-enabled biomanufacturing facilities. By fostering public-private partnerships, the Plan aims to anchor advanced biologics production on U.S. soil, securing supply chains for critical therapeutics.

Workforce Development for AI-Driven Biopharma

Deploying AI across R&D and manufacturing requires a workforce fluent in data science, machine learning, and bioengineering. The Plan establishes:

  • An AI Biopharma Training Hub under the Department of Labor, partnering with community colleges and research institutions to deliver certificates in AI-augmented drug development workflows.
  • Tax guidance clarifying that AI upskilling programs qualify for educational assistance credits, incentivising employers to cover training costs for lab technicians, process engineers, and regulatory specialists.
  • A Rapid Retraining Pilot to transition professionals displaced by automation into high-value roles such as AI model validation, clinical data analysis, and regulatory compliance.

By bridging the skills gap, these initiatives ensure that U.S. biopharma firms have the talent pipeline needed to fully leverage AI’s potential.

Advancing AI Interpretability and Regulatory Trust

One barrier to regulatory acceptance of AI in drug approvals is model opacity. To address this, the Plan calls for:

  • A DARPA-led program to develop interpretable AI architectures that provide mechanistic insights into molecular design decisions.
  • Joint AI hackathons where academia and industry teams stress-test models for robustness, adversarial resilience, and explainability.
  • NIST-sponsored guidelines on AI evaluation metrics tailored to biopharmaceutical use cases, facilitating transparent communication with the FDA.

These efforts aim to build trust between AI developers and regulators, smoothing the path for AI-designed therapeutics to enter clinical trials.

Global Leadership and Export Controls

The U.S. seeks to establish its AI-driven biopharma innovations as the global gold standard. The Plan includes:

  • Export of American AI Platforms for drug discovery to allied nations, combined with joint research initiatives.
  • Tightening of Export Controls on high-performance compute hardware crucial for training large biological foundation models, preventing potential misuse by adversarial states.

By balancing open collaboration with strategic controls, the Plan ensures American biopharma AI remains both influential and secure on the world stage.

Conclusion

America’s AI Action Plan offers a robust blueprint for transforming the pharmaceutical and biopharmaceutical industries. Through investments in automated labs, data infrastructure, biosecurity, manufacturing modernization, workforce training, and interpretability research, it accelerates the journey from molecule to medicine. By anchoring these advances in rigorous safety and regulatory frameworks, the United States is poised to lead a new era of AI-enabled therapeutics—delivering faster cures, lowering costs, and safeguarding public health on a global scale.
Never miss a pharmaceutical headline

The pharmaceutical industry moves fast – stay on top of it with our must - read briefings.

  • The top pharma and life sciences stories, straight to your inbox
  • The biggest news, features, interviews, and analysis
  • Dedicated coverage of the key developments driving the global pharmaceutical sector

Subscribe

- Never miss a story with notifications

- Gain full access to our premium content

- Browse free from any location or device.

Media Packs

Expand Your Reach With Our Customized Solutions Empowering Your Campaigns To Maximize Your Reach & Drive Real Results!

– Access the Media Pack Now

– Book a Conference Call

– Leave Message for Us to Get Back

Latest stories

Related stories

Billerud Introduces New Pharmaceutical Packaging Option

Billerud has officially introduced PharmaKraft, a specialized fibre-based packaging...

AstraZeneca Advances AI Drug Discovery Deal with insitro

AstraZeneca has inked a billion-dollar AI drug discovery partnership...

Managing Quality and Regulatory Risks in Pharmaceutical Technology Transfer

Technology transfer moves more than manufacturing instructions. It moves process knowledge, supplier dependencies, labeling requirements, packaging specifications, and the evidence needed to demonstrate control. For medical device manufacturers, including those supporting drug-device combination products, these interfaces can become recurring sources of compliance gaps during site transitions, acquisitions, and remediation. The central problem is often not a missing procedure. It is a quality system that allows connected activities to change independently. A label revision may be approved while packaging artwork, supplier instructions, or production-system data still reflect an obsolete version. Staying inspection-ready requires manufacturers to manage these dependencies as one controlled system. Why Compliance Gaps Keep Recurring Findings are treated as documentation defects Revising a procedure can resolve an immediate discrepancy without changing the behavior that caused it. If unclear decision rights, weak escalation, or ineffective training remain, the same failure may surface elsewhere. Corrective and preventive action should distinguish the observed finding from its underlying cause and establish how effectiveness will be demonstrated. Ownership is divided across connected functions Quality, Regulatory Affairs, labeling, packaging engineering, operations, suppliers, and digital-system owners often maintain separate workflows. Each team may complete its assigned task while an interface remains uncontrolled. For example, changing an instruction for use can affect packaging configuration, translations, technical documentation, inventory disposition, and release controls. Transitions expose unresolved weaknesses Acquisitions and site transfers can introduce conflicting procedures, incomplete knowledge transfer, or different assumptions about validated processes. Regulatory changes add further demands. When these events coincide with CAPA or change-control backlogs, teams may prioritize administrative closure over risk reduction. Without a coordinated impact assessment, local fixes can create new gaps downstream. Establish the Regulatory Baseline Inspection readiness starts with knowing which requirements apply to the product, activity, and market. Pharmaceutical, device, and combination-product obligations are not interchangeable; transfer planning should identify the applicable requirements rather than assume one quality framework covers every operation. According to FDA’s Quality Management System Regulation overview and FAQs, the QMSR became effective on February 2, 2026, incorporates ISO 13485:2016 by reference, and introduced the updated device inspection process under Compliance Program 7382.850. ISO 13485 certification does not exempt a manufacturer from FDA inspection or guarantee compliance with FDA requirements. For devices marketed in the European Union, Regulation (EU) 2017/745, Annex I, Section 23, provides the primary requirements for information supplied with devices, including labels and instructions for use. Teams should assess the applicable provisions alongside relevant language, product-specific, and market requirements. A labeling review should therefore evaluate required content and consistency with the device’s intended purpose and supporting documentation—not merely artwork accuracy. How to Identify the Real Gaps Begin with a cross-functional assessment that follows actual product and information flows. Select representative transfers, label changes, supplier changes, and CAPAs, then trace each from initiation through implementation and effectiveness review. Map dependencies: Identify affected specifications, suppliers, packaging components, systems, validation activities, and training. Compare records with practice: Check whether approved procedures match what operators and reviewers actually do. Test traceability: Follow a requirement through technical documentation, labeling, production instructions, and release evidence. Examine recurrence: Look for repeated deviations, reopened actions, overdue changes, and findings shared across sites. Assess capacity: Determine whether qualified owners have enough time and authority to complete corrective work. Prioritize gaps by potential patient or user harm, regulatory impact, likelihood of recurrence, and detectability. Document the rationale; an old overdue item is not automatically more important than a newly identified high-risk weakness. Align Quality, Labeling, Packaging, and Change Control Create a coordinated remediation plan with one accountable owner for each action, named contributors, acceptance criteria, and required evidence. Separate immediate containment from systemic correction. Containment might prevent obsolete labels from being issued; correction might redesign approval and implementation controls so obsolete versions cannot return. Use change control as the integration point. Before approval, evaluate effects on regulatory submissions or notifications, technical documentation, supplier controls, packaging performance, validation, electronic master data, inventory, and training. Record both identified impacts and justified decisions that particular activities are unaffected. During technology transfer, define readiness gates before production or release responsibilities move. These should address the transfer of process knowledge, acceptance criteria, receiving-site capability, required qualification or validation, approved documents, and personnel competence. The appropriate evidence depends on the product and process; a generic checklist cannot replace a documented assessment. Finally, verify implementation where the work happens. Confirm that current instructions are available, suppliers received approved requirements, systems contain correct versions, and affected personnel can perform their tasks. A signed approval is not evidence that every downstream change took effect. Mistakes That Undermine Inspection Readiness Closing CAPAs on task completion alone: Effectiveness checks should test whether the underlying failure has been controlled. Updating labels in isolation: Assess packaging, supplier, system, documentation, and inventory consequences. Counting closures instead of evidence: Dashboards need defensible records behind their status indicators. ...

State of API & Peptide CDMO Capacity

Pharmaceutical companies planning peptide or complex small-molecule launches are...

Subscribe

- Never miss a story with notifications

- Gain full access to our premium content

- Browse free from any location or device.

Media Packs

Expand Your Reach With Our Customized Solutions Empowering Your Campaigns To Maximize Your Reach & Drive Real Results!

– Access theMedia Pack Now

– Book a Conference Call

– Leave Message for Us to Get Back

Translate »